Research
Predictive Quantum Chemistry for Excited States and Strong Correlation in Open-Shell Systems
Open-shell molecules and materials—including transition-metal complexes, organic radicals, and defect-based quantum systems—exhibit rich photophysical, catalytic, magnetic, and quantum functionalities. Their theoretical description is challenging because strong correlation, near-degenerate excited states, spin–orbit coupling, and nuclear motion often act simultaneously.
Our group develops spin-adapted time-dependent density functional theory, interacting-reference many-body perturbation theory, and low-complexity correlated wavefunction methods that connect electronic structure with quantitative spectroscopy and dynamics. The three programs below share one goal: making reliable first-principles predictions possible for chemically realistic open-shell systems.
Our central question: How can we develop predictive and scalable electronic-structure theories that connect excited and strongly correlated states in realistic open-shell systems to measurable spectra, dynamics, and chemical reactivity?
Spin-adapted time-dependent density functional theory
Scientific question. How can we compute spin-pure excited-state potential-energy surfaces and interstate couplings with the accuracy and efficiency needed to predict spin conversion and nonadiabatic dynamics in realistic open-shell molecules?
We develop spin-adapted open-shell time-dependent density functional theory, including X-TDDFT for spin-conserving excitations and XSF-TDA for spin-flip-down excitations. Tensor-coupled excitation manifolds remove the spin contamination that limits conventional unrestricted approaches, while retaining the favorable scaling needed for large molecules.
Our current work focuses on analytic gradients, nonadiabatic couplings, spin–orbit interactions, minimum-energy crossing points, and full-dimensional excited-state dynamics. Target applications include transition-metal emitters, organic radicals, spin-dependent photochemistry and catalysis, and molecular and defect-based quantum systems.
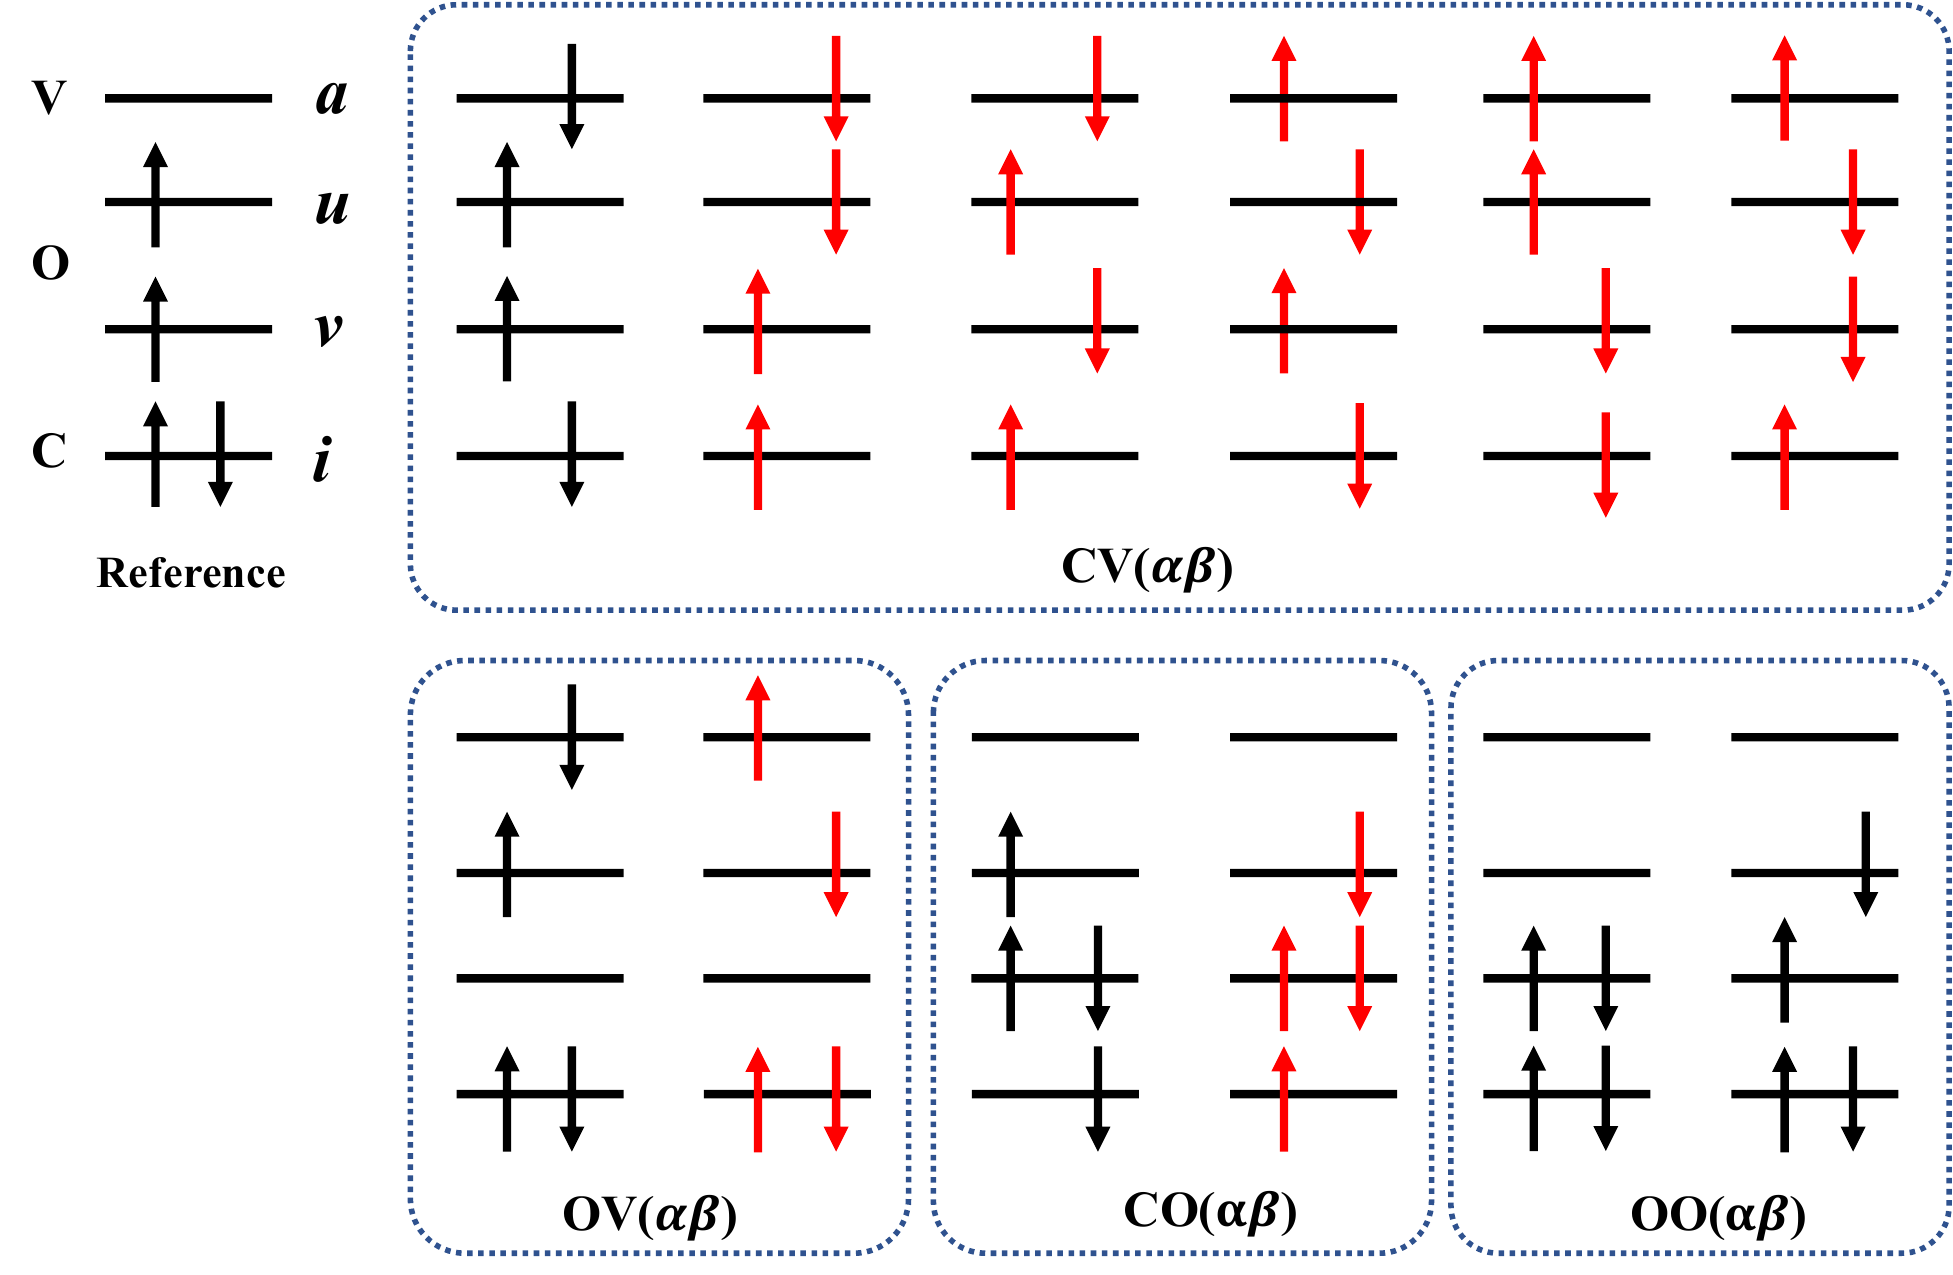
Spin adaptation requires a spin-complete excitation manifold; the missing higher excitations (red) are the source of spin contamination in conventional SF-TD-DFT/SF-TDA.
Selected references
- H. Zhao and Z. Li, “Spin-adapted open-shell time-dependent density functional theory: towards a simple and accurate method for spin-flip-down excitations,” Mol. Phys., e2631735 (2026).
- Z. Li and W. Liu, “First-order nonadiabatic coupling matrix elements between excited states: A Lagrangian formulation at the CIS, RPA, TD-HF, and TD-DFT levels,” J. Chem. Phys. 141, 014110 (2014).
- Z. Li, B. Suo, Y. Zhang, Y. Xiao, and W. Liu, “Combining spin-adapted open-shell TD-DFT with spin–orbit coupling,” Mol. Phys. 111, 3741–3755 (2013).
- Z. Li and W. Liu, “Spin-adapted open-shell random phase approximation and time-dependent density functional theory. I. Theory,” J. Chem. Phys. 133, 064106 (2010).
Interacting-reference many-body perturbation theory
Scientific question. How can many-body perturbation theory be built on an interacting multiconfigurational reference to describe correlation energies, charged excitations, and neutral excitations of strongly correlated systems within a unified diagrammatic framework?
We developed a generalized many-body perturbation theory in which the reference is an interacting multiconfigurational state rather than a single determinant. This construction connects conventional Green’s-function methods in condensed-matter physics with multireference perturbation theory in quantum chemistry.
The first realizations include multireference random-phase approximations in the particle–hole and particle–particle channels. We are extending the same framework toward multireference Green’s-function, GW, and Bethe–Salpeter approaches for quantitative charged and neutral excitations of strongly correlated systems.
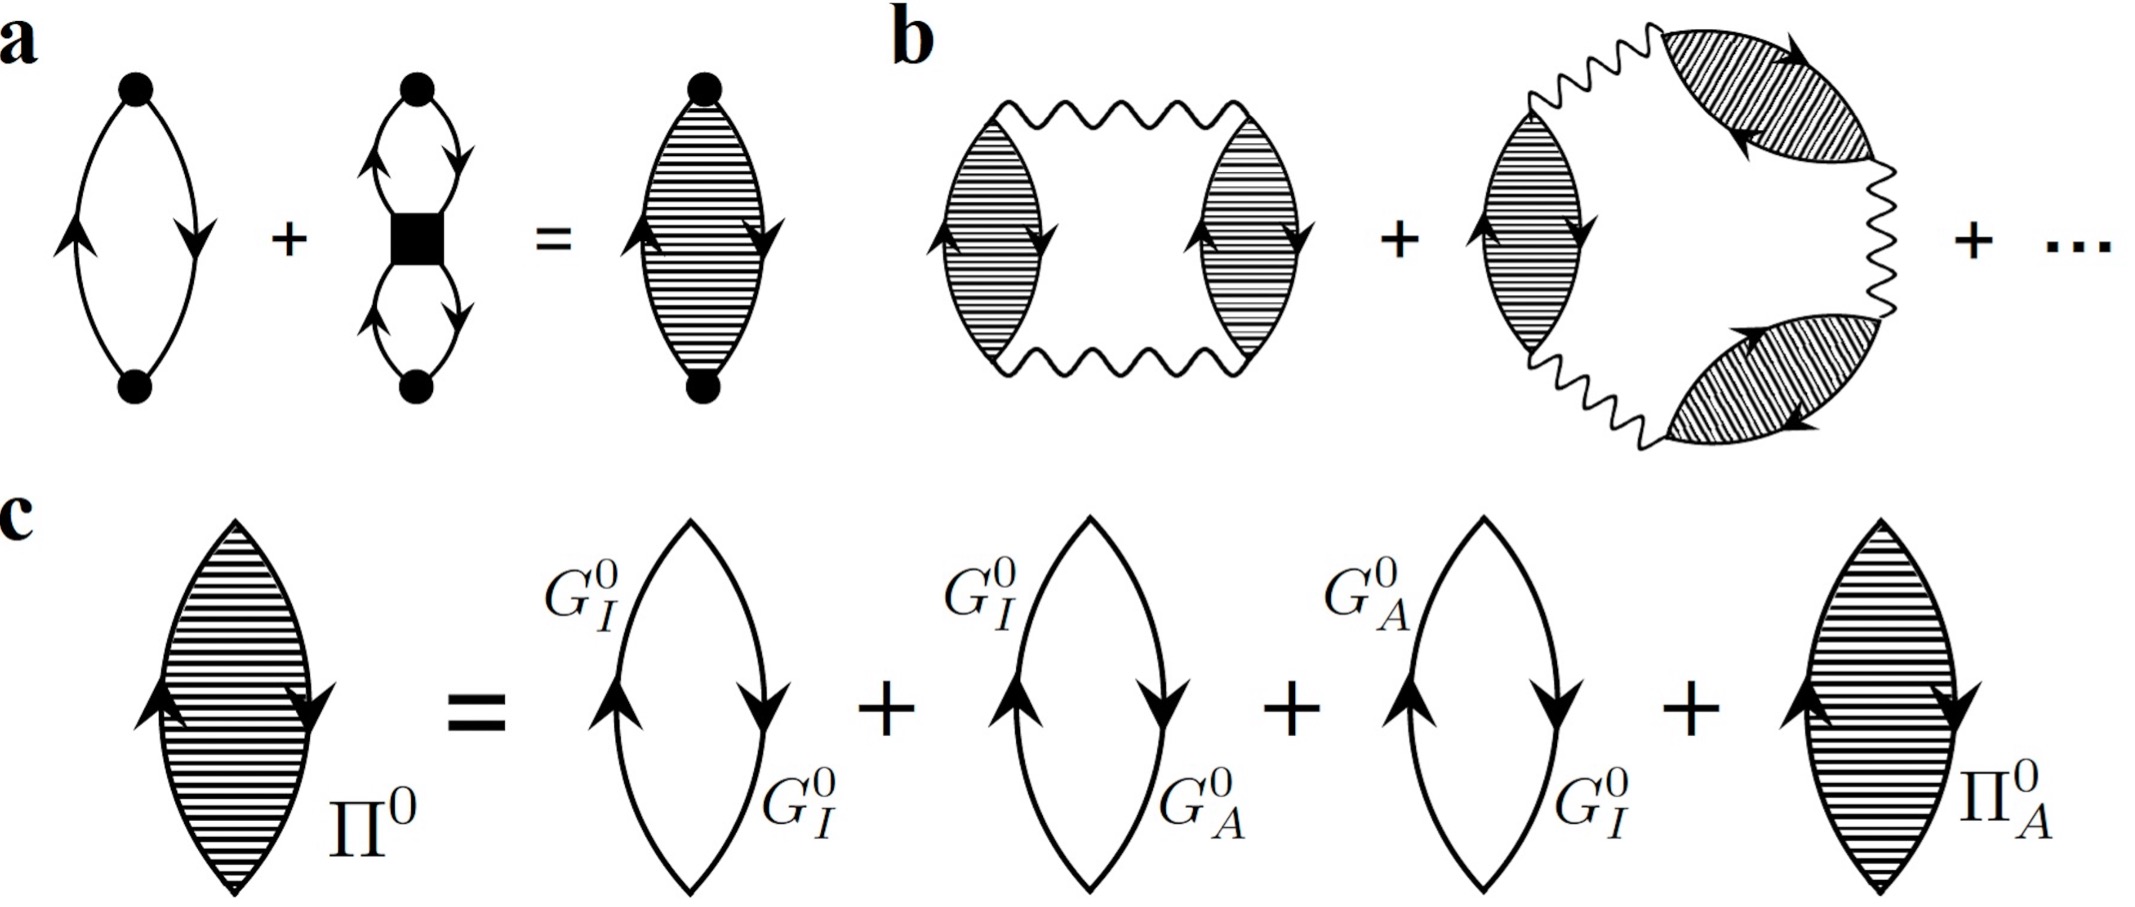
An interacting reference introduces connected Green’s functions and new diagram classes beyond single-reference many-body theory.
Selected references
- Y. Wang, W.-H. Fang, and Z. Li, “Multi-reference GW approximation for strongly correlated molecules,” arXiv:2604.16013 (2026).
- Y. Wang, W.-H. Fang, and Z. Li, “A unified diagrammatic formulation of single-reference and multi-reference random phase approximations: The particle-hole and particle-particle channels,” J. Chem. Phys. 163, 174101 (2025).
- Y. Wang, W.-H. Fang, and Z. Li, “Generalized many-body perturbation theory for the electron correlation energy: Multireference random phase approximation via diagrammatic resummation,” J. Phys. Chem. Lett. 16, 3047–3055 (2025).
- Z. Li, “Stochastic many-body perturbation theory for electron correlation energies,” J. Chem. Phys. 151, 244114 (2019).
Low-complexity correlated wavefunction methods
Scientific question. How can spin symmetry, orbital entanglement, and locality be exploited to construct low-complexity correlated states and reduce the classical and quantum resources required for large-active-space electronic structure and photochemistry?
This program follows two complementary routes—classical many-body methods and chemically informed quantum algorithms. The same structure—spin symmetry, orbital entanglement, locality, and multireference character—guides both routes.
Classical algorithms for strong correlation
We develop tensor-network states and neural-network quantum states to go beyond conventional matrix product states. The goal is to obtain controllable, high-accuracy wavefunctions for active spaces that are inaccessible to conventional multireference methods, especially in polynuclear transition-metal clusters. Our current work combines physically structured wavefunction ansätze with modern GPU and AI-accelerator hardware.
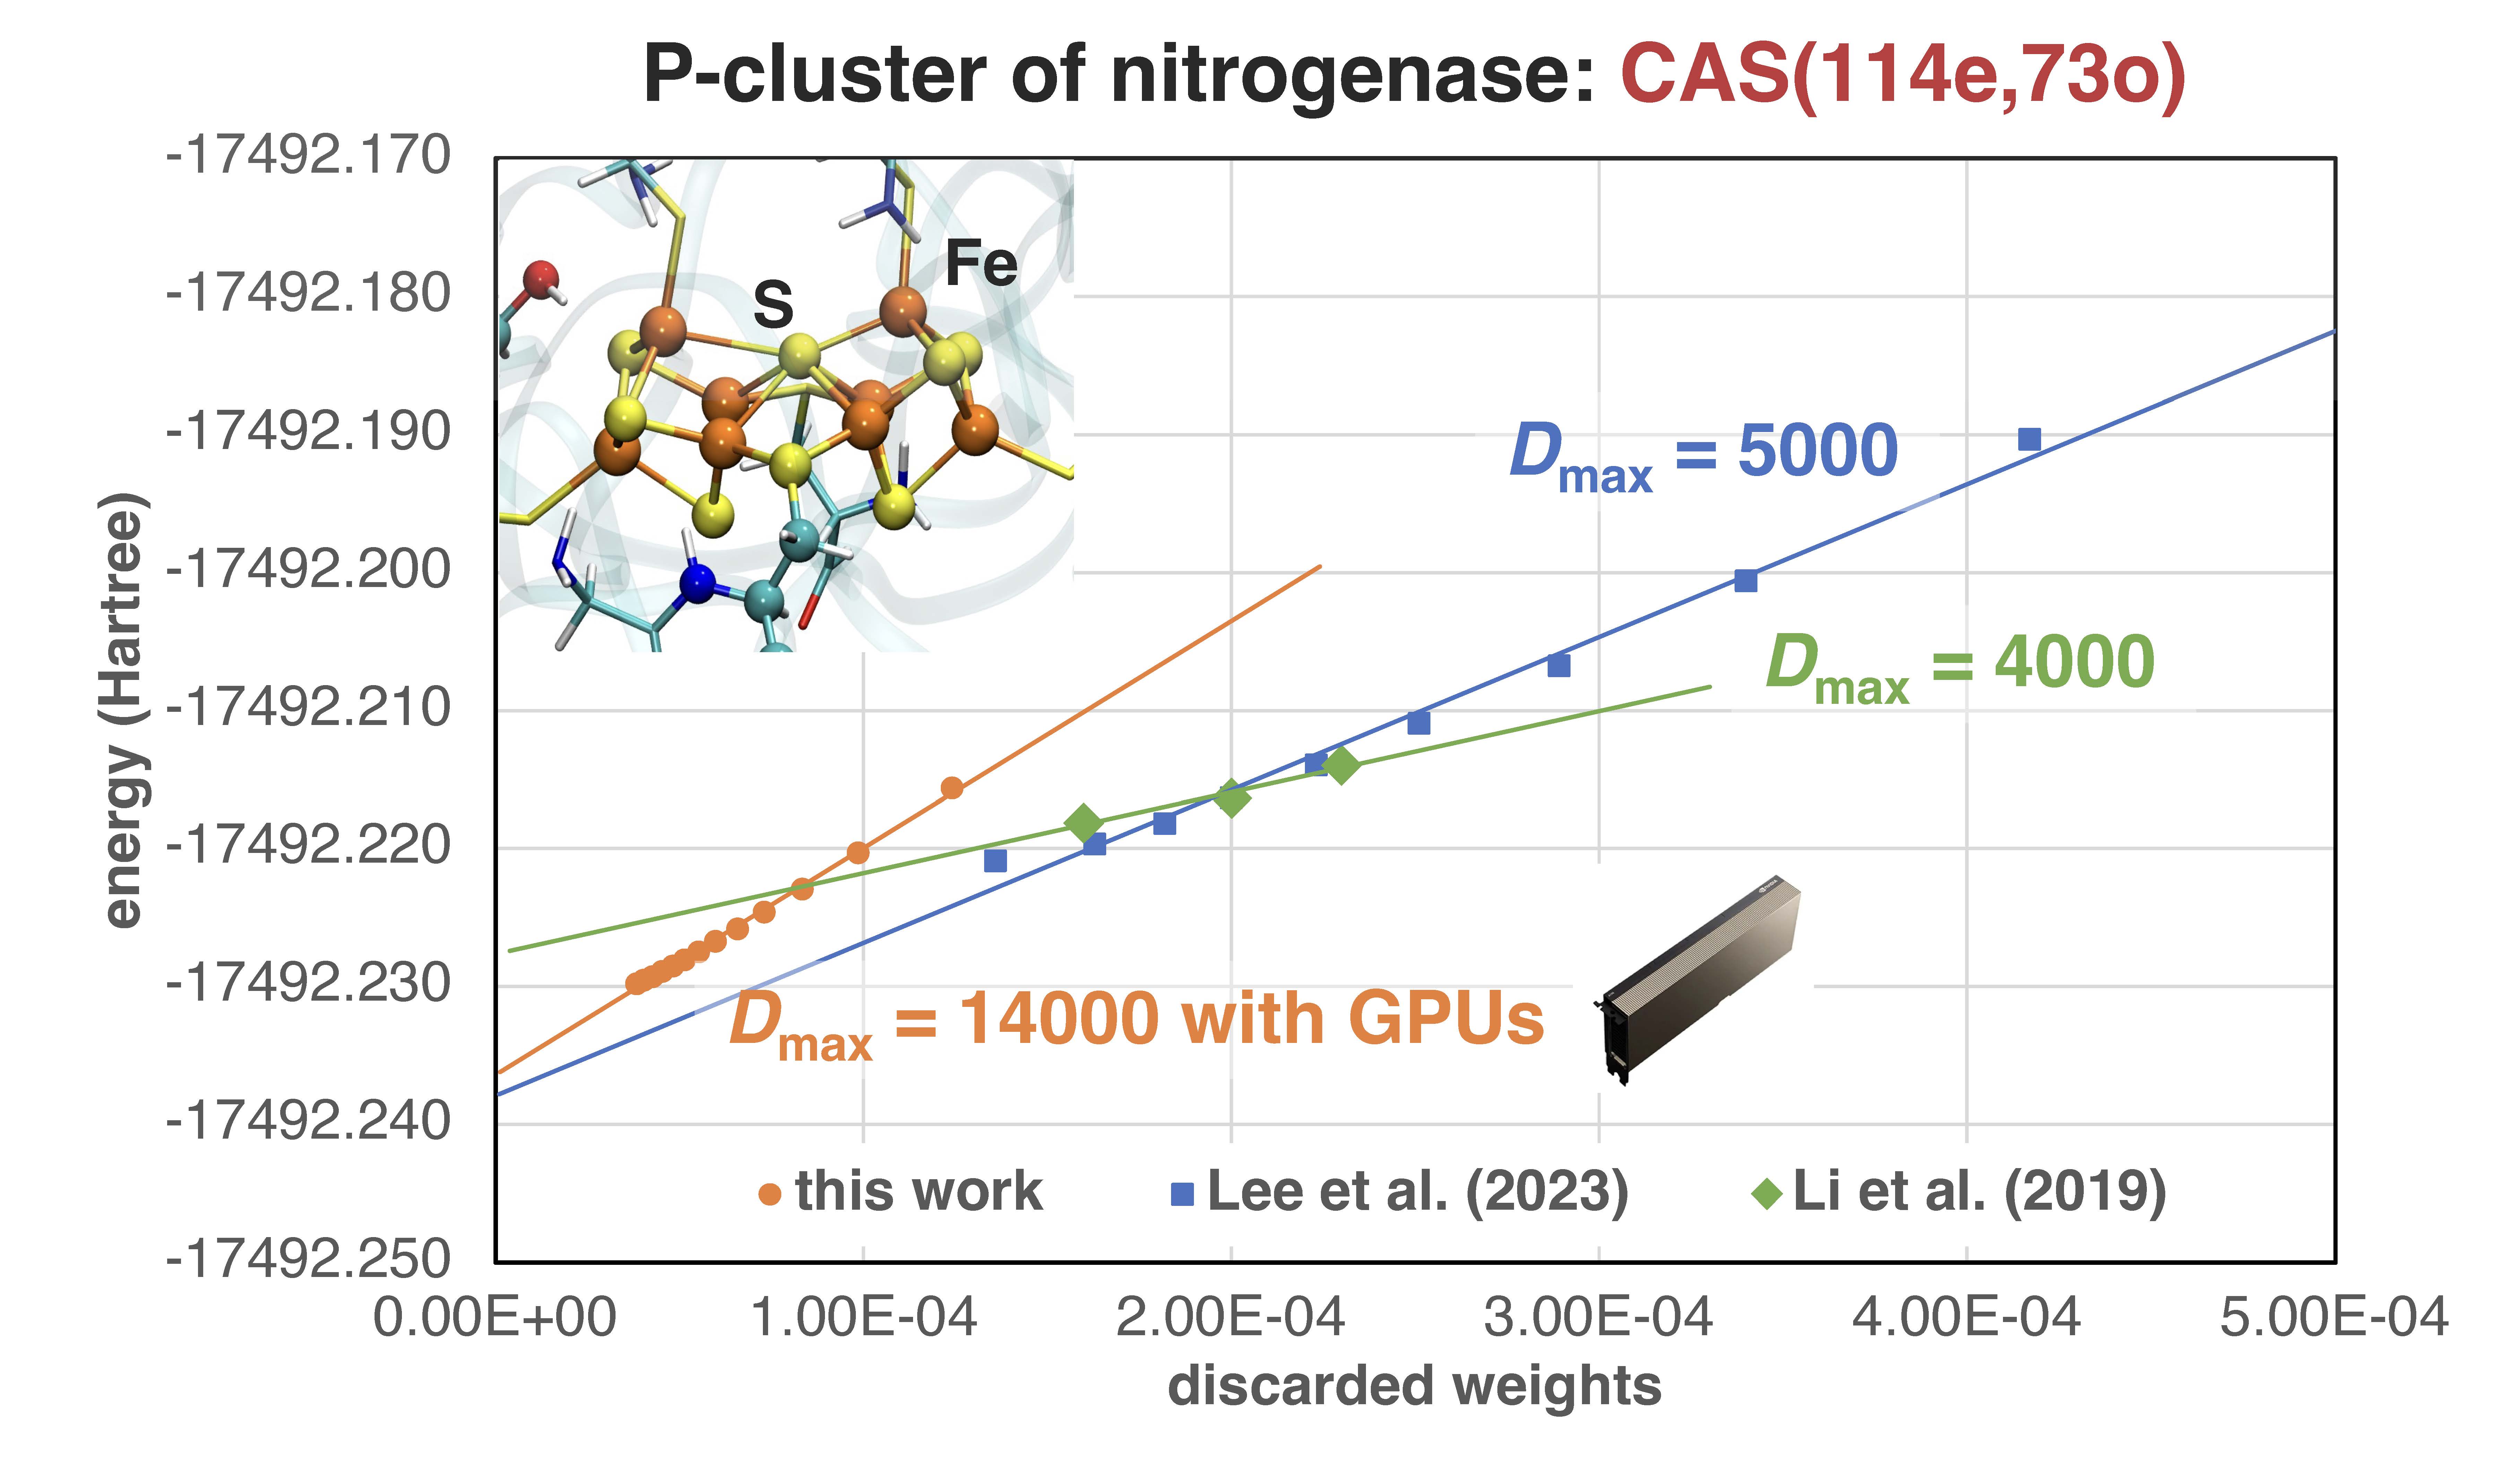
Distributed GPU algorithms make unprecedented active spaces accessible for polynuclear transition-metal clusters.
Selected references
- Y. Li, Z. Wu, B. Zhang, W.-H. Fang, and Z. Li, “Spin-adapted neural network backflow for symmetry-preserving simulations of strongly correlated electrons,” arXiv:2604.06841 (2026).
- Z. Wu, B. Zhang, W.-H. Fang, and Z. Li, “Hybrid tensor network and neural network quantum states for quantum chemistry,” J. Chem. Theory Comput. 21, 10252–10262 (2025).
- C. Xiang, W. Jia, W.-H. Fang, and Z. Li, “Distributed multi-GPU ab initio density matrix renormalization group algorithm with applications to the P-cluster of nitrogenase,” J. Chem. Theory Comput. 20, 775–786 (2024).
Quantum algorithms for electronic structure and photochemistry
We develop quantum algorithms for electronic structure and photochemistry, including efficient state preparation, molecular response properties, and conical intersections. A central theme is to reduce quantum-resource requirements by making maximal use of classical computation. For instance, entanglement-minimized orbitals are constructed using inexpensive classical tensor-network calculations to produce compact initial states, directly connecting our classical many-body expertise with quantum phase estimation for challenging systems such as iron–sulfur clusters.
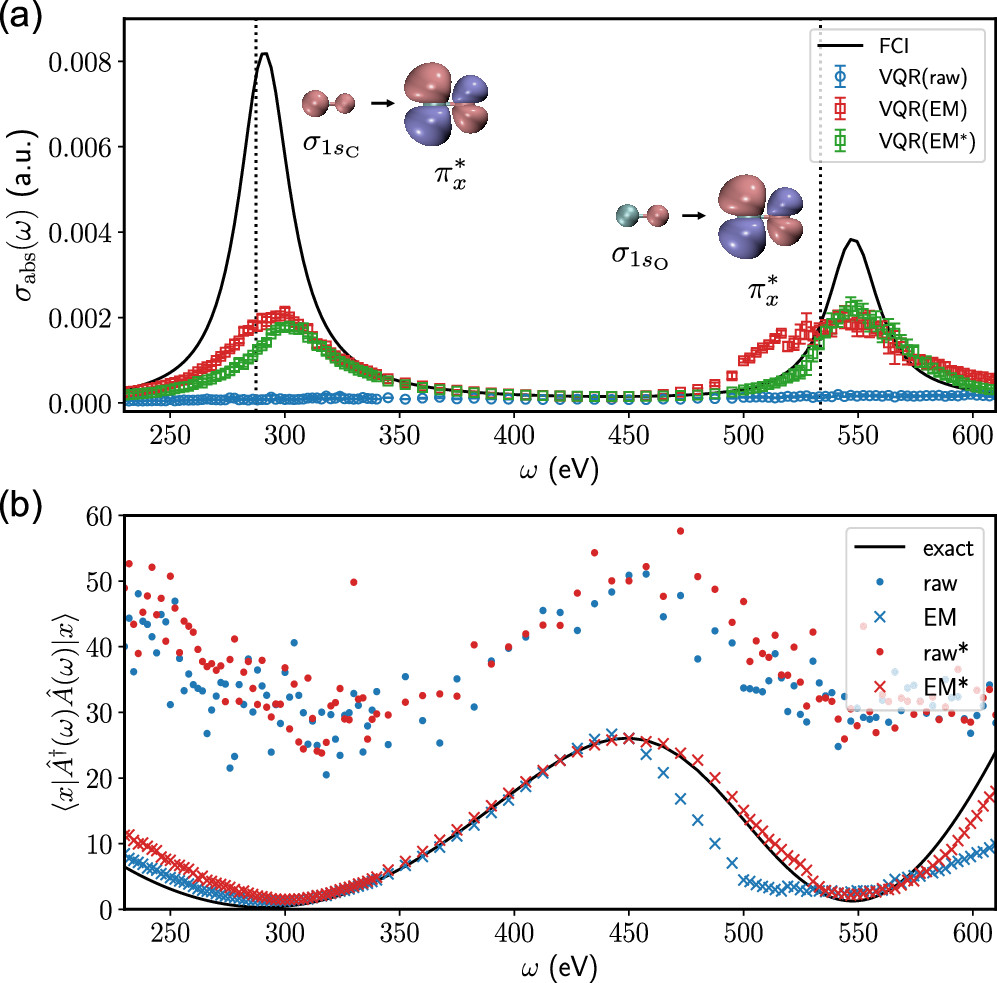
Quantum algorithms connect molecular response, photochemical intersections, and strongly correlated state preparation with experimentally accessible quantum hardware.
Selected references
- Z. Li, “Entanglement-minimized orbitals enable faster quantum simulation of molecules,” Phys. Rev. Lett. 135, 210601 (2025).
- S. Zhao, D. Tang, X. Xiao, R. Wang, Q. Sun, Z. Chen, X. Cai, Z. Li, H. Yu, and W.-H. Fang, “Quantum computation of conical intersections on a programmable superconducting quantum processor,” J. Phys. Chem. Lett. 15, 7244–7253 (2024).
- K. Huang, X. Cai, H. Li, Z.-Y. Ge, R. Hou, H. Li, T. Liu, Y. Shi, C. Chen, D. Zheng, K. Xu, Z.-B. Liu, Z. Li, H. Fan, and W.-H. Fang, “Variational quantum computation of molecular linear response properties on a superconducting quantum processor,” J. Phys. Chem. Lett. 13, 9114–9121 (2022).
Where theory meets chemistry
We focus on chemical problems in which excited states, strong correlation, spin, and nuclear motion are inseparable—and where predictive theory can reveal mechanisms or observables inaccessible to standard approaches.
Spin-dependent photophysics and spectroscopy
- Open-shell photodynamics: predicting spin conversion, nonadiabatic relaxation, and spin-forbidden emission in organic radicals, transition-metal complexes, and other open-shell molecules;
- Quantum-system photophysics: determining how excited-state structure, spin–orbit and vibronic interactions control spin polarization, coherence, optical initialization, and readout in molecular and defect-based systems;
- Multimodal spectroscopy: combining optical, EPR, X-ray, and Mössbauer observables to assign correlated electronic states, distinguish competing mechanisms, and guide molecular design.
Spin-dependent reactivity in biological and materials systems
- Polynuclear transition-metal systems: resolving exchange coupling, dense electronic manifolds, and reactivity in molecular clusters, metalloenzyme active sites, and materials containing multiple correlated open shells;
- Spin-selective catalysis: determining how competing spin surfaces, surface crossings, charge transfer, and nonadiabatic branching control activity and selectivity in catalysis and photocatalysis.
The long-term objective is a connected theoretical stack: scalable classical wavefunctions provide accurate references and compact state representations; quantum algorithms open complementary routes to response properties and state preparation; interacting-reference many-body theory adds systematic dynamical correlation; and spin-adapted response theory turns these electronic states into measurable spectra, dynamics, and chemical mechanisms.
For a complete publication list, see Google Scholar. For current opportunities, see Join Us.